
tophat输出结果junction.bed
tophat输出结果junction.bed
| BED format |
|
BED format provides a flexible way to define the data lines that are displayed in an annotation track. BED lines have three required fields and nine additional optional fields. The number of fields per line must be consistent throughout any single set of data in an annotation track. The order of the optional fields is binding: lower-numbered fields must always be populated if higher-numbered fields are used. If your data set is BED-like, but it is very large and you would like to keep it on your own server, you should use the bigBed data format. The first three required BED fields are:
The 9 additional optional BED fields are:
Example: track name=pairedReads description="Clone Paired Reads" useScore=1 Example: browser position chr7:127471196-127495720 Example: browser position chr7:127471196-127495720 |
tophat输出结果junction.bed中定义的junction就是中间是intron两边是exon的这种序列结构, JUNC00000001包含两个exon和中间的intron
cat junctions.bed
track name=junctions description="TopHat junctions"

test_chromosome 177     400     JUNC00000001    61      +       177     400     255,0,0 2       73,50   0,173

test_chromosome 350     550     JUNC00000002    45      +       350     550     255,0,0 2       50,50   0,150
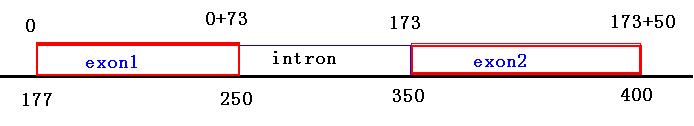
exon1: 177-250(display), 177-249(actual)
intron: 250-350(display), 250-349(actual)
exon2: 350-400(display), 350-399(actual)
chrom(test_chromosome): The name of the chromosome.
这里就是test_chromosome
chromStart(177): The starting position of the feature in the
chromosome or scaffold. The first base in a chromosome is numbered
0.
chromosome上feature(在这里就是junction)的起始位置
chromEnd(400): The ending position of the feature in the chromosome
or scaffold. The chromEnd base is not included in the display of
the feature. For example, the first 100 bases of a chromosome are
defined as chromStart=0, chromEnd=100, and span the bases numbered
0-99.
对应的终止位置
name(JUNC00000001): Defines the name of the BED line.
score(61): A score between 0 and 1000.
strand(+): Defines the strand: either '+' or '-'.
thickStart(177): The starting position at which the feature is
drawn thickly (for example, the start codon in gene
displays).
thickEnd(400): The ending position at which the feature is drawn
thickly (for example, the stop codon in gene displays).
itemRgb(255,0,0): An RGB value of the form R,G,B (e.g. 255,0,0). If
the track line itemRgb attribute is set to "On", this RBG value
will determine the display color of the data contained in this BED
line. NOTE: It is recommended that a simple color scheme (eight
colors or less) be used with this attribute to avoid overwhelming
the color resources of the Genome Browser and your Internet
browser.
blockCount(2): The number of blocks (exons) in the BED line.
外显子个数,2个
blockSizes(73,50): A comma-separated list of the block sizes. The
number of items in this list should correspond to
blockCount.外显子大小,两个数字用逗号隔开
blockStarts(0,173): A comma-separated list of block starts. All of
the blockStart positions should be calculated relative to
chromStart. The number of items in this list should correspond to
blockCount.
外显子起始位置,不从chromosome的起始开始,而是从chromStart开始数,也就是junction的起始位置开始计算,第一个碱基是0
用bed_to_junc转换后得到.juncs其中只保留了junction中intron的部分
cat new_list.juncs
test_chromosome 249     350     +

test_chromosome 399     500     +
<chrom> <left> <right> <+/->
http://genome.ucsc.edu/FAQ/FAQformat.html#format1
http://www.ensembl.org/info/website/upload/bed.html
http://zhongguozhanying.blog.163.com/blog/static/21618709520135484450746/
http://blog.sina.com.cn/s/blog_7cffd14001012rtd.html
https://github.com/WormBase/wormbase-pipeline/blob/master/scripts/align_RNASeq.pl
tophat输出结果junction.bed的更多相关文章
- 使用Tophat+cufflinks分析差异表达
使用Tophat+cufflinks分析差异表达 2017-06-15 19:09:43 522 0 0 使用TopHat+Cufflinks的流程图 序列的比对是RNA分析 ...
- RNA-seq连特异性
RNA-seq连特异性 Oct 15, 2015 The strandness of RNA-seq analysis 前段时间一直在研究关于illumina TrueSeq stranded RNA ...
- tophat的用法
概述:tophat是以bowtie2为核心的一款比对软件. tophat工作分两步: 1.将reads用bowtie比对到参考基因组上. 2.将unmapped-reads打断成更小的fragment ...
- TopHat
What is TopHat? TopHat is a program that aligns RNA-Seq reads to a genome in order to identify exon- ...
- RNA-seq差异表达基因分析之TopHat篇
RNA-seq差异表达基因分析之TopHat篇 发表于2012 年 10 月 23 日 TopHat是基于Bowtie的将RNA-Seq数据mapping到参考基因组上,从而鉴定可变剪切(exon-e ...
- poj 1041(字典序输出欧拉回路)
John's trip Time Limit: 1000MS Memory Limit: 65536K Total Submissions: 8641 Accepted: 2893 Spe ...
- 链终止法|边合成边测序|Bowtie|TopHat|Cufflinks|RPKM|FASTX-Toolkit|fastaQC|基因芯片|桥式扩增|
生物信息学 Sanger采用链终止法进行测序 带有荧光基团的ddXTP+其他四种普通的脱氧核苷酸放入同一个培养皿中,例如带有荧光基团的ddATP+普通的脱氧核苷酸A.T.C.G放入同一个培养皿,以此类 ...
- mapreduce多文件输出的两方法
mapreduce多文件输出的两方法 package duogemap; import java.io.IOException; import org.apache.hadoop.conf ...
- Android Studio 多个编译环境配置 多渠道打包 APK输出配置
看完这篇你学到什么: 熟悉gradle的构建配置 熟悉代码构建环境的目录结构,你知道的不仅仅是只有src/main 开发.生成环境等等环境可以任意切换打包 多渠道打包 APK输出文件配置 需求 一般我 ...
随机推荐
- [DevExpress]ChartControl之滚动条示例
关键代码: /// <summary> /// 设置ChartControl滚动条[默认X,Y轴都出现] /// </summary> /// <param name=& ...
- Linux 服务器如何禁止 ping 以及开启 ping
Linux 默认是允许 ping 响应的,也就是说 ping 是开启的,但 ping 有可能是网络攻击的开始之处,所以关闭 ping 可以提高服务器的安全系数.系统是否允许 ping 由2个因素决定的 ...
- C++ list 类学习笔记
双向循环链表list list是双向循环链表,,每一个元素都知道前面一个元素和后面一个元素.在STL中,list和vector一样,是两个常被使用的容器.和vector不一样的是,list不支持对元素 ...
- shell 实现类似php的require_once函数
config.sh #/bin/bash require_once() { #File the true path ,To prevent a symbolic link local realpath ...
- 监听EditText
0.得到焦点的时候,作一些处理 public class AbcActivity extends Activity implements OnFocusChangeListener{ @Overrid ...
- MySQL创建数据库[保存mojo格式的数据库]已经常用的utf8格式数据库
一.创建最新编码utf8mb4格式的库: CREATE DATABASE IF NOT EXISTS yourdbname CHARACTER SET utf8mb4 COLLATE utf8mb4_ ...
- PHP+MYSQL 出现乱码的解决方法
PHP+MYSQL 出现乱码的解决方法 使用PHP+MYSQL时遇到过字符乱问题,解决方法: 在mysql_connect后面加一句SET NAMES UTF8,即可使得UTF8的数据库消除乱码,对于 ...
- Java类加载器加载类顺序
java ClassLoader的学习 java是一门解释执行的语言,由开发人员编写好的java源文件先编译成字节码文件.class形式,然后由java虚拟机(JVM)解释执 行,.class字节码文 ...
- easy ui 下拉框绑定数据select控件
easy ui 中的下拉框控件叫做select,具体代码如下: html代码:①.这是一个公司等级的下拉框 <tr> <td>公司等级:</td> <td&g ...
- 【BZOJ 1189】[HNOI2007]紧急疏散evacuate
Description 发生了火警,所有人员需要紧急疏散!假设每个房间是一个N M的矩形区域.每个格子如果是'.',那么表示这是一块空地:如果是'X',那么表示这是一面墙,如果是'D',那么表示这是一 ...